-
Glass Fused To Steel Tanks (896)
-
Stainless Steel Tanks (434)
-
Fusion Bonded Epoxy Tanks (652)
-
Galvanized Steel Tanks (321)
-
Aluminum Dome Roofs (1336)
-
Wastewater Storage Tanks (280)
-
Welded Steel Tanks (457)
-
Pressure Vessels (426)
-
Anaerobic Digester (202)
-
Industrial Water Tanks (349)
-
Glass Lined Steel Tanks (180)
-
Bolted Steel Tanks (389)
-
Sludge Storage Tank (115)
-
Biogas Storage Tank (173)
-
Leachate Storage Tanks (133)
-
Agricultural Water Storage Tanks (179)
-
Fire Water Tank (166)
-
Grain Storage Silos (146)
-
Biogas Projects (424)
-
Wastewater Treatment Projects (1712)
-
Double Membrane Roof (223)
|
What Is a Pharmaceutical Reactor? GMP Design, Materials & Applications
| Place of Origin: | China |
| Brand Name: | Center Enamel |
| Certification: | ISO 9001,CE, NSF/ANSI 61, WRAS, ISO 28765, LFGB, BSCI, ISO 45001 |
| Minimum Order Quantity: | 1 Sets |
| Price: | 2000 |
| Delivery Time: | 2 months |
| Payment Terms: | L/C,T/T |
| Supply Ability: | 200 sets / days |
|
Detail Information |
|||
Product Description
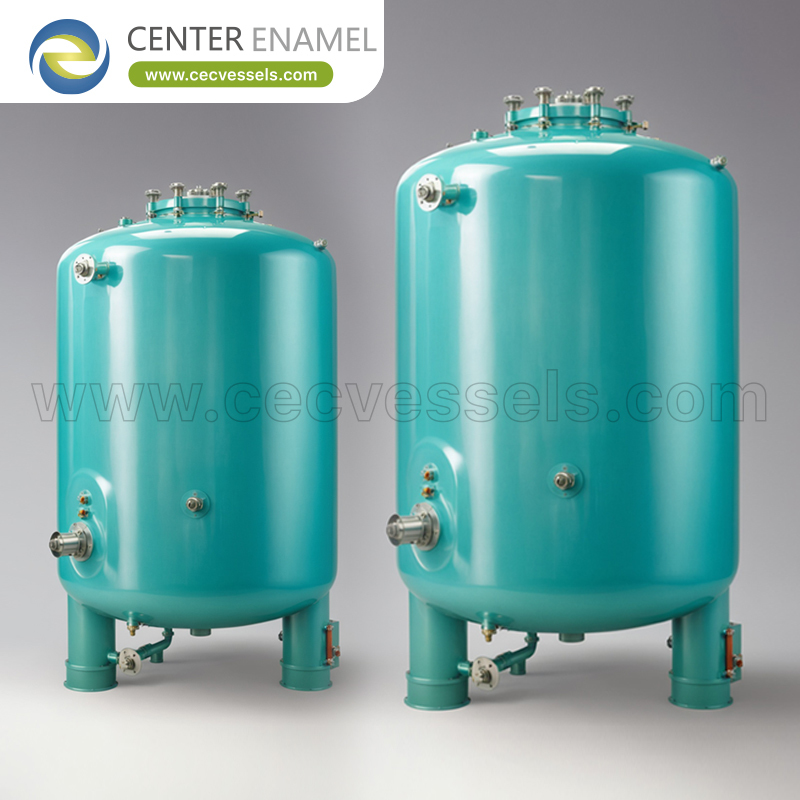
What Is a Pharmaceutical Reactor? GMP Design, Materials & Applications
Answering the core question: What is a pharmaceutical reactor, and how does it meet GMP and product-safety requirements? A pharmaceutical reactor is a cGMP-compliant vessel for manufacturing active pharmaceutical ingredients (APIs) and biologics, built to ASME-BPE with electropolished product-contact surfaces (Ra <= 0.5 µm, often 0.25-0.38 µm), full drainability (3D rule), and validated CIP/SIP (121°C for 30 min, Fo >= 15; sterile assurance SAL 10-6). It operates under 21 CFR Part 210/211 (and for biologics ICH Q5A), with PAT and QbD controlling critical quality attributes and 21 CFR Part 11 / ALCOA+ data integrity for batch records. Materials are 316L (or higher for corrosive steps), passivated per ASTM A967, with extractables/leachables assessed per USP <665>. The reactor is IQ/OQ/PQ qualified and designed so every surface is cleanable and inspectable.
1. Core GMP Principles of Pharmaceutical Reactors
Pharma reactors are defined by quality systems, not just hardware. Three GMP pillars govern design:
- **Surface, Cleanability and Sterility:** Product-contact surfaces are electropolished to Ra <= 0.5 µm (biologics 0.25-0.38 µm) and passivated (ASTM A967) to form a passive, non-adherent chromium oxide layer. All lines are 3D drainable (no dead legs) to allow validated CIP (0.5-1.5% NaOH at 60-80°C, 60-90 min) and SIP (121°C x 30 min, Fo >= 15). Aseptic reactors achieve SAL 10-6; bioburden on cleaned surfaces is < 1 CFU/25 cm2, verified by swab and media-fill.
- **Quality by Design, PAT and Data Integrity:** Under ICH Q8/Q11 (QbD), the reactor is designed around critical process parameters (CPP) and critical quality attributes (CQA) controlled by PAT (in-situ Raman/NIR, off-gas, biomass). Batch records follow 21 CFR Part 11 with ALCOA+ (attributable, legible, contemporaneous, original, accurate, plus complete, consistent, enduring, available) data integrity. This enables real-time release and continuous process verification rather than end-of-batch testing.
- **Material Compatibility and Extractables:** 316L is standard; higher alloys (904L, Hastelloy, zirconium) are used for corrosive API steps. For single-use or polymeric parts, extractables/leachables are qualified per USP <661>/<665> (and <1665>) with a safety threshold against dose. Metal leaching, catalyst residues, and cleaning-agent carryover are bounded by cleaning validation (10 ppm / 1/1000 dose criteria) so cross-contamination between products is prevented.
2. Major Pharmaceutical Reactor Types
Pharma reactors split by molecule class. Three types dominate API and biologic manufacture:
- **API Synthesis Reactor (chemical):** A jacketed 316L STR (0.5-20 m3) for multi-step small-molecule API synthesis under cGMP. It carries accountable-material handling, inert blanketing, and full traceability; clean between products via validated CIP to 10 ppm residue limit. Often Hastelloy/zirconium lined for halogenated or acidic steps. BPCS enforces master formula; deviations are investigated per deviation system.
- **Biopharma Bioreactor (biological):** A 316L/SUB STR (50-20,000 L) for cell culture or microbial fermentation of biologics (mAb, vaccine, recombinant protein). ASME-BPE, CIP/SIP, kLa 50-200 h-1, DO/pH control, PAT for viable cell density and titer. Single-use variants (50-2,000 L) cut cleaning validation. Sterility SAL 10-6 and viral safety per ICH Q5A are designed in.
- **Sterile / Aseptic Reactor (injectable):** A reactor for sterile API or formulation feeding aseptic filling, built to 3-A/EHEDG and sterile-design principles with SIP and aseptic transfer. Surface Ra <= 0.38 µm, no dead legs, HEPA/sterile-filter protected. Used where the product is terminally sterilized or aseptically processed for injectables, with the highest cleaning and monitoring rigor.
Pharmaceutical Reactor Type Comparison Matrix
Type |
Surface / Code |
Sterility |
Primary Product |
API Synthesis Reactor |
316L, ASME-BPE, Ra 0.5 |
CIP-validated |
Small-molecule API |
Biopharma Bioreactor |
316L/SUB, BPE, SIP |
SAL 10-6 |
mAb, vaccine, protein |
Sterile Aseptic Reactor |
Ra 0.38, 3-A/EHEDG |
Aseptic transfer |
Injectable API |
Frequently Asked Questions (FAQ)
Q: What surface finish is required for a GMP pharmaceutical reactor?
A: Product-contact surfaces must be electropolished to Ra <= 0.5 µm (biologics typically 0.25-0.38 µm) and passivated per ASTM A967, with all welds full-penetration and crevice-free and all drains 3D drainable. This ensures validated cleanability, prevents biofilm, and supports SAL 10-6 sterility for aseptic service. Roughness is documented in IQ/OQ and re-verified after major repairs.
Q: What is the difference between cGMP for API vs biologics reactors?
A: Both follow 21 CFR 210/211, but biologics add ICH Q5A (viral safety), cell-bank control, and sterile-process rigor (SAL 10-6, media fills) plus ASME-BPE. Small-molecule API reactors emphasize cleaning validation (cross-contamination 10 ppm), accountable material, and chemical traceability. Biologic reactors emphasize aseptic operation, cell-culture control, and extractables from single-use films.
Q: How is data integrity assured on a pharma reactor?
A: The DCS/historian enforces 21 CFR Part 11 and ALCOA+ (attributable, legible, contemporaneous, original, accurate, complete, consistent, enduring, available): electronic signatures, audit trails, time-stamped batch records, and access control. No manual data overwrite; deviations are logged. This lets regulators trust the electronic batch record for release and continuous verification.
Q: What cleaning validation limits apply between products?
A: Cleaning validation proves residue below 10 ppm of previous product in the next, or 1/1000 of the minimum daily dose (the stricter), with bioburden < 1 CFU/25 cm2 and no detectable cleaning-agent carryover. Swab and rinse samples verify it; the CIP recipe and 3D-drainable design make the limit achievable and reproducible across campaigns.